genomics

The Vertebrate Genomes Project Phase I: A global reference genome resource
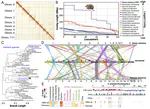
The Vertebrate Genomes Project (VGP) aims to produce complete and near-error-free reference genomes for all ∼70,000 extant vertebrate species1. Organized in four phases, it progressively targets all vertebrate orders, families, genera, and eventually all species. Here we present the completion of VGP Phase I, delivering reference genomes for ∼95% of vertebrate orders, along with additional lineages within those orders, totaling 816 species and 1.6 trillion base pairs of main haplotype sequence. These genomes were assembled and annotated over an 8-year period (2018-2026) of rapid advances in genome sequencing, assembly, and annotation methods2–4, alongside the growth of associated consortium initiatives and international collaborations5–9. They represent some of the highest-quality vertebrate genomes currently available, and most have become the primary reference for their respective species in public databases. Comparative analyses across a subset of 579 species when we reached a threshold of 85% of orders allowed us to reconstruct the genome of the last common ancestor of all vertebrates 500 million years ago, identify diverse modes of sex chromosome evolution, reveal clade-specific three-dimensional genome architecture, discover methylated epigenetic landscapes across vertebrates, and provide a framework for studying gene and pseudogene evolution, immune loci, cancer-associated genes, and other trait-associated loci. Approximately a quarter of this subset are listed as Vulnerable to Critically Endangered by the IUCN Red List of Threatened Species, and have enabled more advanced genomic investigations of extinction risk. VGP Phase I delivers a reference backbone for vertebrate genomics, enabling discoveries that would otherwise remain out of reach across evolution, conservation, and medicine.
Exon Targeted Retrieval and Classification Toolbox (ExTRaCT): a gene search pipeline to find APOBEC3 Z-domains in novel bat genomes
Motivation Several computation gene search tools exist to identify and annotate an ever-growing body of newly sequenced genomes of different species. Many annotation tools, however, fall short when the target species diverges from well-studied model organisms, and when searching for short genes with multiple copies. Results We have developed the Exon Targeted Retrieval and Classification Toolbox, ExTRaCT, an automated pipeline to identify any gene exon with conserved structure in novel species genome assemblies. In the use cases presented here, we applied our search tool to 102 bat genomes to find APOBEC3 gene family members. We show that our homolog search algorithm is efficient (run time average of 5 hours for over 100 genomes), works well with reference sequences distantly related to the target (1 out of 498 misclassifications, 0 false positives and 2 false negatives), and is easy to use. As genomic sequencing becomes faster and more accessible, ExTRaCT has downstream applications in phylogenetic, biochemical and genomic studies. It is a simple computational tool that provides a solution to target gene identification, requiring neither whole-genome-assembly annotations, nor prior knowledge of closely related species.
Genomic comparisons and the adaptive basis of brain size plasticity and chromosomal instability in the Eurasian common shrew
Sorex araneus, the Eurasian common shrew, has seasonal brain size plasticity (Dehnel’s phenomenon) and abundant intraspecific chromosomal rearrangements, but genomic contributions to these traits remain unknown. We couple a chromosome-scale genome assembly with seasonal brain transcriptomes to discover relationships between molecular changes and both traits. Positively selected genes enriched the Fanconi anemia DNA repair pathway, which prevents the accumulation of chromosomal aberrations, and is likely involved in chromosomal rearrangements (FANCI, FAAP100). Genes involved in neurogenesis show either signatures of positive selection (PCDHA6), seasonal differential expression in the cortex and hippocampus (Notch signaling), or both (SOX9), suggesting a role for cellular proliferation in seasonal brain shrinkage and regrowth. Both positive selection and evolutionary upregulation in the shrew hypothalamus of VEGFA and SPHK2 indicate adaptations in hypothalamic metabolic homeostasis have evolved together with Dehnel’s phenomenon. These findings reveal genomic changes central to the evolution of both chromosomal instability and cyclical patterns in brain gene expression that characterizes mammalian brain size plasticity.
Bat genomes illuminate adaptations to viral tolerance and disease resistance
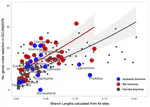
Zoonoses are infectious diseases transmitted from animals to humans. Compared to other mammalian orders, bats are suggested to harbor more zoonotic viruses(Olival et al. 2017). Infections in bats are largely asymptomatic(Schlottau et al. 2020; Guito et al. 2021), suggesting limited tissue-damaging inflammation and immunopathology. To investigate the genomic basis of disease resistance, the Bat1K project generated reference-quality genomes of ten bat species, including potential viral reservoirs. A systematic analysis covering 115 mammalian genomes revealed that signatures of selection in immune genes are more prevalent in bats compared with other mammalian orders. We found an excess of immune gene adaptations in the ancestral chiropteran branch and in many descending bat lineages, highlighting viral entry and detection factors, and regulators of antiviral and inflammatory responses. ISG15, an antiviral gene contributing to hyperinflammation during COVID-19(Perng and Lenschow 2018; Munnur et al. 2021), exhibits key residue changes in rhinolophid and hipposiderid bats. Cellular infection experiments show species-specific antiviral differences and an essential role offor protein conjugation in antiviral function of bat ISG15, separate from its role in secretion and inflammation in humans. Furthermore, in contrast to human ISG15, ISG15 of most rhinolophid and hipposiderid bats has strong anti-SARS-CoV-2 activity. Our work reveals molecular mechanisms contributing to viral tolerance and disease resistance in bats.
Newly assembled pocket gopher genomes can facilitate conservation management of biodiversity
Texas exhibits one of the richest levels of pocket gopher diversity in the United States. Three genera (Cratogeomys, Geomys, and Thomomys) and 11 species are found in Texas. It is not surprising given the diversity of the Texas landscape (ecoregions, life zones, substrates, and vegetation) that these species are further subdivided into 29 subspecies in Texas alone. Pocket gopher distributions are determined by availability of suitable soil types and therefore often occur in small, isolated populations. For some taxa, limited distribution and ultimately small deme sizes result in populations that may require attention from a regulatory and management perspective. For many Texas pocket gopher subspecies, insufficient information exists to make sound recommendations relative to conservation status and needs despite decades of research collecting and evaluating data based on morphometrics, distributions and habitat preferences, karyotypes, allozymes, and mitochondrial DNA. As such, there is precedent for elevating pocket gopher subspecies to species after evaluation of available data, as well as subsuming subspecies into a broader taxonomic group. We used genomic techniques to identify genetically defined operational taxonomic units (OTUs) of pocket gophers to improve knowledge and understanding of pocket gopher distributions within the state. Using tens of thousands of single nucleotide polymorphisms, we determined the number of OTUs in each genus to be 5 for Thomomys bottae subspecies, 8 for Geomys species, and 5 for Cratogeomys castanops subspecies in Texas. In general, these data agree with current taxonomic hypotheses regarding Geomys and C. castanops; however, many T. bottae groups present similar genetic patterns that do not merit subspecies status based on these data, suggesting a more conservative classification of T. bottae in Texas and southeastern New Mexico that could facilitate conservation efforts, should they be necessary.
The molecular basis of viral tolerance in bats
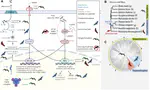
The ability of bats to act as natural reservoir hosts of zoonotic viruses has been attributed to derived aspects of their innate immune systems. In particular, bats appear to detect and respond to pathogens differently compared to humans, allowing them to tolerate viruses that are harmful to other species. Studies to date have identified several lineage-specific mechanisms responsible for dampened immune and inflammatory responses in bats; however, these have mainly focused on a few putative reservoir species and their relatives, representing ~1% of extant bat species diversity. We will conduct the first large-scale study of bat immune adaptations by screening >150 genes in 300 species from across the bat clade, spanning >60 million years of evolution. We will apply sequence capture to obtain orthologues, and perform analyses of selection, parallelism and functional impact to identify compelling putative molecular adaptations. To assess the impact of lineage-specific putative adaptations on immune responses, we will then conduct functional assays on bat and human cells exposed to viruses. For this we will focus on the proteins STING, NLRP3 and MyD88 (which encompass central effector pathways for activating IFN, inflammasome and NFkB, respectively), in each case using CRISPR/Cas9 to build transgenic cell lines that differ with respect to key residues. Finally, we will examine whether the presence of impactful molecular adaptations in these and other loci can explain known variation in bat-virus interactions.
Human and bats genome robustness under COSMIC mutational signatures
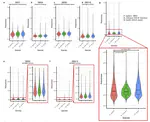
Carcinogenesis is an evolutionary process, and mutations can fix the selected phenotypes in selective microenvironments. Both normal and neoplastic cells are robust to the mutational stressors in the microenvironment to the extent that secure their fitness. To test the robustness of genes under a range of mutagens, we developed a sequential mutation simulator, Sinabro, to simulate single base substitution under a given mutational process. Then, we developed a pipeline to measure the robustness of genes and cells under those mutagenesis processes. We discovered significant human genome robustness to the APOBEC mutational signature SBS2, which is associated with viral defense mechanisms and is implicated in cancer. Robustness evaluations across over 70,000 sequences against 41 signatures showed higher resilience under signatures predominantly causing C-to-T (G-to-A) mutations. Principal component analysis indicates the GC content at the codon’s wobble position significantly influences robustness, with increased resilience noted under transition mutations compared to transversions. Then, we tested our results in bats at extremes of the lifespan-to-mass relationship and found the long-lived bat is more robust to APOBEC than the short-lived one. By revealing APOBEC as the prime driver of robustness in the human (and other mammalian) genome, this work bolsters the key potential role of APOBECs in carcinogenesis, as well as evolved countermeasures to this innate mutagenic process. It also provides the baseline of the human and bat genome robustness under mutational processes associated with cancer.